Custom Specialty Aromatic Derivative CDMO Solutions for Complex Synthesis Challenges
2026-10-31
Every complex synthesis route eventually hits a wall—poor selectivity, stubborn impurities, or a step that refuses to scale. At DSL Chemicals, we don't treat these as dead ends; we treat them as starting points. Our custom specialty aromatic derivative CDMO solutions are built around the messy, unpredictable realities of advanced chemistry, turning process bottlenecks into reliable, scalable pathways. From route scouting to cGMP manufacturing, we partner with teams who need more than a vendor—they need a problem-solving extension of their own lab. Here's how we approach the synthesis challenges most CROs avoid.
Where Standard Routes Stall We Step In
Standard routes are built for predictable volumes, clean data, and the smooth middle of the bell curve. The moment an order carries an unusual dimension, a delivery window shifts mid-transit, or a supplier falls silent on a Friday afternoon, the carefully mapped process starts to fray. That is exactly where we pick up the thread—not by forcing the exception back into the template, but by treating the outlier as the main event.
Our team steps in when the handoff between systems, carriers, or departments leaves a gap that no dashboard can flag. We work the phones, the back channels, and the loading docks until the shipment moves again. It is not about adding more steps to the same route; it is about building a short, direct path around the blockage while keeping every stakeholder in the loop.
The result is not a patched-together fix that barely holds. It is a deliberate intervention that restores momentum, protects the delivery promise, and leaves behind a quieter, more resilient route for the next time the standard playbook stalls.
Custom Aromatic Cores Built Around Your Molecule
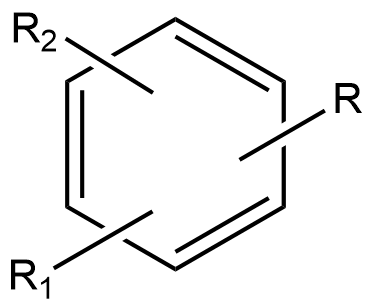
Most catalog building blocks force your target into a shape that already exists. We take the opposite route: the aromatic core is assembled around the specific geometry, electronic demands, and steric constraints of your molecule. Whether the goal is to tune a binding pocket, shift a reduction potential, or lock a conformation, the ring system is drawn and synthesized to fit the problem rather than bending the problem to fit a stock scaffold.
That means you are not limited to phenyl, pyridyl, or the usual fused systems. Mixed heteroaromatic cores, unusual substitution patterns, nonplanar twists, and ring fusions that are absent from commercial libraries can be built from the ground up. Each design is routed through viable synthetic chemistry before it reaches the bench, so what looks like an exotic core on paper remains practical to prepare at the scale you actually need.
Because the core is built around your molecule, late-stage surprises tend to surface earlier. Solubility, metabolic soft spots, crystallization behavior, and synthetic accessibility are considered as part of the core design rather than treated as downstream fixes. The result is not another aromatic fragment from a list—it is a structural decision made for your compound.
Scaling Complex Derivatives Without Losing Selectivity
In synthetic chemistry, the jump from milligram-scale reactions to multi-kilogram production often breaks the very selectivity that made a route attractive in the first place. Complex derivatives—molecules with multiple reactive centers, delicate stereochemistry, or late-stage functional groups—tend to misbehave when heat transfer slows, mixing becomes imperfect, or local concentration gradients form. The challenge is not merely reproducing a yield; it is preserving the precise regiochemical or stereochemical outcome that was carefully engineered at small scale.
One practical path is to redesign the reaction around scale, rather than forcing the scaled vessel to mimic the flask. This might mean switching from batch to continuous flow, where residence time and thermal control stay sharp even at high throughput. Alternatively, protecting-group strategies or catalytic systems can be tuned so that selectivity arises from intrinsic kinetics rather than from fragile mixing patterns. The key is to identify which parameter actually drives the desired pathway—temperature, stoichiometry, concentration, or catalyst loading—and then hold that parameter constant as the reactor grows.
Successful scale-up of complex derivatives rarely comes from a single heroic change. It emerges from iterative stress testing: deliberately perturbing mixing, heat removal, and feed rates at intermediate scale to map out where selectivity collapses. Once those boundaries are known, the process can be engineered with comfortable margins. The result is a synthesis that delivers the same molecular precision at ton scale as it did on the bench, without sacrificing the selectivity that made the molecule worth making in the first place.
Solubility and Stability Fixes for Stubborn Intermediates
Getting a stubborn intermediate to dissolve can feel like coaxing a rock into solution. The usual suspects—switching from hexanes to ether, warming the flask, or adding a drop of DMF—rarely cut it when the molecule has a dense ring system or a zwitterionic core. Salt formation often works better than expected: a quick counterion swap to a tosylate or mesylate can turn an oily mess into a crystalline solid that precipitates cleanly. For intermediates that refuse to behave, an in situ quench or a temporary protecting group on a polar handle sometimes loosens the lattice enough to keep the reaction moving.
Stability is a different beast. Intermediates that decompose on silica, oxidize in air, or dimerize within hours need more than a good solvent trick. Storing them as a frozen solution in toluene or THF under argon can buy a few extra days, but if you need to isolate them, lyophilization from aqueous acetonitrile or a cold ether trituration often beats column chromatography. For truly touchy cases, generating the intermediate and feeding it directly into the next step—without ever isolating—turns the stability problem into a scheduling problem, which is usually easier to solve.
A CDMO That Actually Enjoys Steric and Electronic Puzzles
Most CDMOs treat hindered couplings or electron-poor building blocks as a necessary evil—something to push through with standard conditions and hope for the best. We take a different view. Steric and electronic puzzles are where our chemists actually lean in, mapping out the orbital mismatches, rotating around congested bonds, and tuning ligands until a supposedly impossible cross-coupling starts behaving. It is not about brute force; it is about enjoying the intellectual challenge hidden inside each stubborn intermediate.
That mindset shows up in the way we scout catalysts, select bases, and adjust solvent polarity for each project. Instead of forcing a generic Pd cycle onto a molecule that clearly has other ideas, we study what the substrate is telling us—whether it wants a bulky N-heterocyclic carbene, a softer halide, or a completely different metal altogether. The result is a route that respects the molecule's personality rather than fighting it.
For clients, this means fewer rescue batches, shorter timelines, and a partner who is genuinely curious about difficult chemistry. Whether you are facing a tri-ortho-substituted aryl ring or a heterocycle that refuses to participate in standard aminations, you get a team that treats the obstacle as the most interesting part of the day. That is the difference between a CDMO that merely tolerates complexity and one that looks forward to it.
Process Chemistry for Molecules That Refuse to Behave
Some molecules seem to have a personality problem. You draw a clean route on the whiteboard, run the first step at small scale, and it works. Then the pilot batch turns the same intermediate into tar, or the product refuses to crystallize until you seed it with material from a different polymorph. Process chemistry for these stubborn compounds is less about optimizing yield and more about negotiating with a reactant that does not follow the usual rules.
A classic offender is the heteroaromatic ring that insists on protonating where you least expect it, turning a simple alkylation into a messy mixture. Instead of forcing conditions, process chemists often change the order of bond formation. Move the fragile coupling to a point where the offending nitrogen is already tied up in a stable intermediate, and the reaction suddenly behaves. The same logic applies to protecting groups: choose one that falls off under mildly acidic conditions if the rest of the molecule hates strong acid.
Temperature control becomes a negotiation too. A molecule that tolerates heating for hours may not survive a hot filtration, while another decomposes at room temperature but is perfectly stable at -20°C. Flow reactors, controlled quenches, and non-traditional solvents all earn their place in the toolbox. The satisfaction comes when a process that once failed at every turn finally holds up across multiple batches, not because the molecule changed, but because the chemistry learned to accommodate it.
FAQ
A standard supplier sells catalog compounds, but a CDMO focused on aromatic derivatives takes your synthetic route and owns the process—scaling it, troubleshooting impurities, and handling the regulatory documentation. The specialty part means we're comfortable with awkward ring functionalization, selective halogenation, or making low-volume intermediates that bigger players won't touch.
We lean on in-house screening of directing groups and catalysts rather than guessing. For example, if you need a meta-substituted arene that typical electrophilic substitution won't give, we'll test masked directing strategies or transition-metal-catalyzed C–H activation early, so you don't waste months on a dead-end route.
We routinely work from 100 milligram quantities for early medicinal chemistry exploration up to multi-kilogram batches for preclinical and early clinical supply. The scaling isn't just about bigger reactors—it's about re-optimizing exotherms, mixing, and workups so the selectivity holds when you go from a 250 mL flask to a 50 L vessel.
Yes, but we treat those as normal engineering problems rather than exotic exceptions. We have dedicated Schlenk lines, gloveboxes, and contained filtration setups, and our staff does routine training on handling boronic ester intermediates, azides, and other energetic or toxic species. The key is designing the process so the hazardous step happens in a closed system from day one.
We don't just run a standard LCMS and call it done. We use a combination of high-resolution mass spec, 2D NMR (like HSQC and HMBC), and chiral or achiral SFC when needed. For close isomers, we often develop a custom HPLC method with a stationary phase that actually separates your specific pair, rather than relying on generic C18 columns.
We write up the process as if we're handing it to someone who has never seen the chemistry—including all the failure points, impurities we chased down, and the exact quality of raw materials that mattered. We also offer a shadow-run period where your team can observe batches and ask questions before taking over.
Rough ideas are actually more common than fully developed routes. We can start with a literature precedent, a patent scheme, or even just a target structure and run feasibility studies. Our chemists will propose two or three orthogonal routes and test them in parallel, so you get a data-driven recommendation rather than a single biased opinion.
We use separate project codes, restricted access to electronic lab notebooks, and physical segregation for client materials. Our contracts default to client ownership of any process improvements or new derivatives we generate, and we're happy to sign CDAs or joint development agreements tailored to your legal team's needs.
Conclusion
When established synthetic pathways collapse under the weight of steric hindrance, fragile functionality, or aggressive coupling conditions, our team treats the dead end as a starting point. We design aromatic cores around the specific demands of your molecule—not the other way around—so electron-rich, electron-poor, or highly substituted rings become an advantage rather than a liability. Scaling these derivatives from milligram to kilogram is handled with the same obsession for selectivity: chiral centers hold, regiochemistry stays locked, and impurities that typically emerge at pilot scale are engineered out of the process long before they appear.
Insoluble intermediates that turn every workup into a fight and unstable species that decompose on contact with silica get reformulated through salt screening, co-crystal approaches, or tailored protecting group strategies. We genuinely enjoy the puzzles that make other CDMOs walk away—twisted biaryls, atropisomeric scaffolds, nitrogen-rich heterocycles with unpredictable electronics. The result is a process chemistry function built for molecules that refuse to behave, delivering robust, repeatable routes without sanding off the complexity that makes your target valuable.
Contact Us
Contact Person: Wei Zhang
Email: [email protected]
Tel/WhatsApp: 862163529955
Website: https://www.dslchem.com